|
Search Term: " rimehylglycine "
The Ultimate Guide to Cellular Longevity: NAD+, Methylation, and Senolytics   
Date:
September 10, 2026 10:57 AM
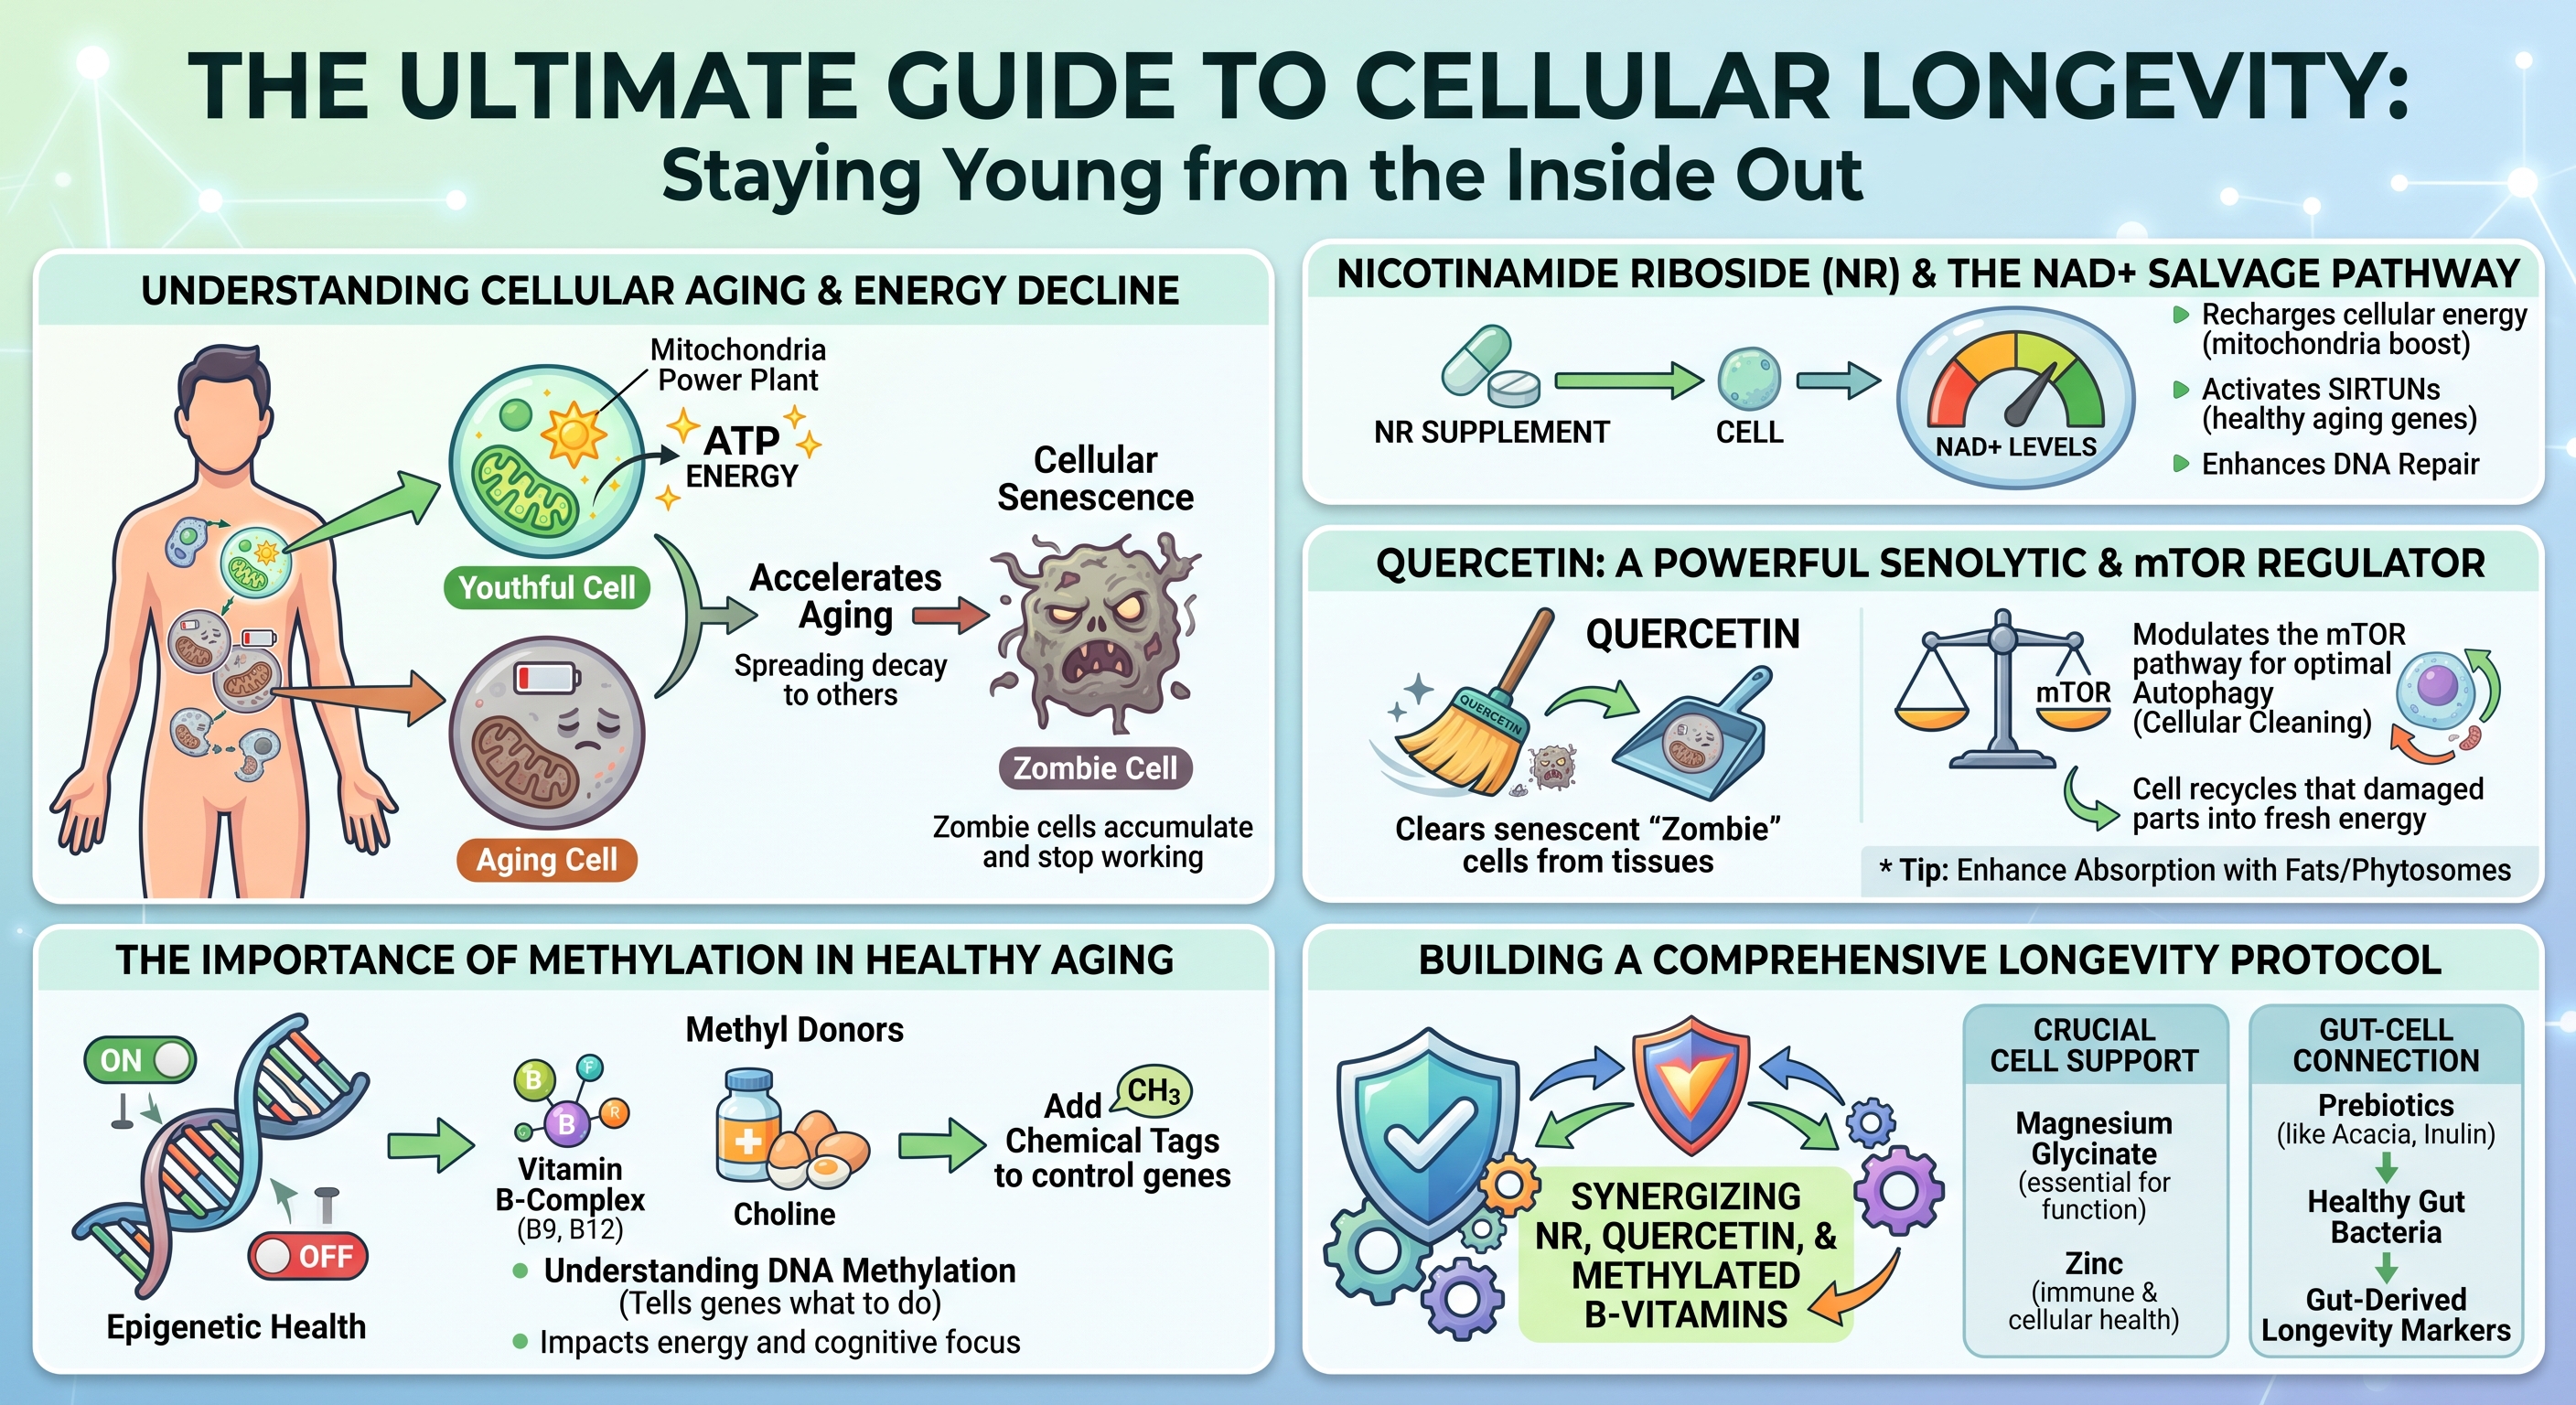
Introduction: Understanding Cellular Aging and Energy DeclineBiological aging represents a progressive decline in cellular maintenance, structural repair, and energy generation. Over decades, tissues experience an attrition of functional reserves, compromised stress resilience, and persistent low-grade systemic inflammation. At the cellular scale, biological degeneration is driven by a failure to generate bioenergetic fuel, repair genetic code, and clear metabolic waste.Cellular aging is characterized by interconnected biological disruptions known as the hallmarks of aging. These encompass genomic instability, epigenetic alterations, mitochondrial decay, loss of proteostasis, and cellular senescence. Rather than operating as isolated occurrences, these phenomena establish a self-reinforcing degenerative cycle: declining cellular power generation impairs enzymatic genetic repair, promoting the accumulation of damaged cells that enter irreversible growth arrest and poison surrounding healthy tissues. Mitigating cellular aging requires examining how microscopic bioenergetic pathways deteriorate and evaluating how targeted nutritional and biochemical interventions can restore cellular homeostasis. The Role of Mitochondria and ATP ProductionEvery biological function - from muscular contraction to continuous DNA replication - depends on adenosine triphosphate (ATP), the primary biochemical energy currency of living systems. Cells produce the vast majority of this energy within mitochondria through oxidative phosphorylation. Within these specialized organelles, metabolic intermediates derived from dietary carbohydrates and lipids donate high-energy electrons to the electron transport chain. The flow of these electrons across protein complexes establishes an electrochemical proton gradient across the inner mitochondrial membrane, driving ATP synthase to manufacture ATP.A youthful cell functions like an efficient municipal power grid, dynamically matching energetic demands with immediate ATP output. However, as biological aging progresses, mitochondrial efficiency declines. The electron transport chain becomes structurally leaky, inadvertently shedding electrons that react with ambient molecular oxygen to produce reactive oxygen species (ROS). While regulated levels of ROS participate in vital intracellular signaling, chronic excess induces widespread oxidative stress. Mitochondria are exceptionally vulnerable to this oxidative burden because they carry their own circular genetic material, known as mitochondrial DNA (mtDNA). Unlike nuclear DNA, mtDNA lacks the protective shielding of histone proteins and possesses rudimentary repair systems. As a result, mtDNA sustains cumulative oxidative damage, encoding increasingly defective electron transport chain proteins. This dynamic generates a bioenergetic deficit: degraded mitochondria synthesize progressively less ATP while emitting greater volumes of damaging free radicals. Deprived of optimal ATP reserves, cells lack the energy necessary to drive vital enzymatic repair cascades, accelerating structural degeneration and functional exhaustion. How Cellular Senescence Accelerates the Aging ProcessWhen healthy cells confront critical physiological damage - such as severe telomere attrition, persistent DNA double-strand breaks, or oxidative stress - they activate protective cell cycle arrest pathways governed primarily by the p53/p21^CIP1 and p16^INK4a/Rb molecular checkpoints. This defensive shutdown, termed cellular senescence, permanently prevents the replication of potentially premalignant or mutated cells.Senescent cells, colloquially known as "zombie cells," enter a state of permanent growth arrest while actively resisting programmed cell death (apoptosis). Over time, these cells accumulate within adipose depots, skeletal muscle, the vascular endothelium, and major organs, largely because immune surveillance and clearance pathways simultaneously lose functional efficiency. The systemic danger of senescent cells stems from their secretome. Rather than remaining biologically inert, senescent cells develop a hyperactive secretory state termed the Senescence-Associated Secretory Phenotype (SASP). The SASP is a destructive mixture of pro-inflammatory cytokines, chemokines, extracellular matrix-degrading matrix metalloproteinases (MMPs), and reactive oxygen species. Through this toxic secretome, even a small burden of senescent cells can impair whole-tissue architecture. SASP factors degrade surrounding structural proteins, induce insulin resistance in neighboring metabolic cells, and biochemically force adjacent healthy cells into secondary senescence. This persistent paracrine signaling fuels chronic, sterile, low-grade systemic inflammation, termed "inflammaging," which accelerates systemic tissue degeneration and elevates susceptibility to degenerative age-related pathologies. Nicotinamide Riboside (NR) and the NAD+ Salvage PathwayThe Biochemistry of NAD+ Depletion Over TimeNicotinamide adenine dinucleotide (NAD+) is an indispensable coenzyme present in every living cell. NAD+ fulfills a dual biological mandate: it serves as a central redox cofactor that shuttles electrons between cellular metabolic reactions, and it functions as an obligatory consumable substrate for regulatory enzymes that preserve cellular viability. In its redox capacity, NAD+ accepts electrons to form NADH during glycolysis, the tricarboxylic acid (TCA) cycle, and fatty acid beta-oxidation, subsequently donating those electrons to Complex I of the respiratory chain to power ATP synthesis.
The primary enzymatic driver of age-related NAD+ destruction is CD38, a membrane-bound glycohydrolase expressed on immune cells that is upregulated in response to chronic SASP exposure. Concurrently, lifelong genotoxic damage causes persistent activation of Poly(ADP-ribose) polymerase 1 (PARP-1), an enzyme that cleaves the glycosidic bonds of NAD+ to assemble branched poly(ADP-ribose) chains at DNA lesion sites. Because PARP-1 consumes NAD+ without directly recycling the molecule, chronic DNA damage depletes intracellular NAD+ pools, impairing bioenergetics and limiting sirtuin activity. How NR Efficiently Boosts Cellular NAD+ LevelsThe mammalian body maintains its NAD+ supply through three distinct biosynthetic routes: the de novo pathway from dietary L-tryptophan, the Preiss-Handler pathway from nicotinic acid (niacin), and the NAD+ Salvage Pathway. The de novo pathway requires substantial energy expenditure, consuming roughly sixty milligrams of dietary tryptophan to yield a single milligram of NAD+. The Preiss-Handler pathway, while effective, can induce cutaneous prostaglandin-mediated flushing at therapeutic intakes. Consequently, the salvage pathway serves as the primary mechanism for maintaining intracellular NAD+ pools.The salvage pathway recycles the breakdown product nicotinamide (NAM), which is released whenever NAD+-consuming enzymes execute their functions. Under normal conditions, cells convert free nicotinamide into nicotinamide mononucleotide (NMN) via the rate-limiting enzyme nicotinamide phosphoribosyltransferase (NAMPT), after which NMN adenylyltransferases (NMNAT1–3) complete the conversion into NAD+. However, NAMPT expression declines with advancing age, chronic inflammation, and metabolic stress, limiting the recycling capacity of the cell. Nicotinamide Riboside (NR) is a naturally occurring pyridine nucleoside that bypasses this enzymatic bottleneck. Upon cellular entry via equilibrative nucleoside transporters, NR is directly phosphorylated into NMN by nicotinamide riboside kinases (NRK1 and NRK2) using a single molecule of ATP. Because the NRK pathway remains intact and robust across the lifespan, NR provides an efficient alternative entry point into the NAD+ salvage cascade. Clinical evaluations in humans confirm the safety, bioavailability, and pharmacokinetics of oral NR supplementation. Randomized, double-blind, placebo-controlled trials reveal that oral NR chloride produces dose-dependent increases in steady-state whole blood NAD+ concentrations. Dosing regimens of 100 mg, 300 mg, and 1,000 mg daily elevate blood NAD+ levels by approximately 22%, 51%, and up to 142%, respectively, within two weeks of administration, maintaining these elevations throughout continuous use. High-resolution metabolomic analyses also demonstrate parallel elevations in nicotinic acid adenine dinucleotide (NAAD), establishing it as a reliable biomarker of active intracellular NAD+ synthesis without hepatic or systemic toxicity. Sirtuin Activation and DNA Repair MechanismsReplenishing intracellular NAD+ supports functions beyond mitochondrial ATP generation. NAD+ functions as an obligatory cofactor for sirtuins (SIRT1 through SIRT7), a family of class III histone and non-histone protein deacetylases that regulate stress resilience, metabolic homeostasis, and cell survival. Sirtuins couple the removal of acetyl groups from target lysine residues to the stoichiometric cleavage of NAD+, producing nicotinamide and O-acetyl-ADP-ribose. In states of NAD+ deficiency, sirtuin enzymes remain inactive regardless of cellular demand.In the nucleus, SIRT1 coordinates defense against cellular decline. When activated by restored NAD+ levels, SIRT1 deacetylates peroxisome proliferator-activated receptor-gamma coactivator 1-alpha (PGC-1a), the master transcriptional coactivator of mitochondrial biogenesis. This deacetylation stimulates mitochondrial replication and assembly, expanding functional respiratory capacity. Concurrently, SIRT1 deacetylates the p65 subunit of nuclear factor-kappa B (NF-kB), suppressing the transcription of pro-inflammatory cytokines. In the mitochondria, SIRT3 utilizes NAD+ to deacetylate metabolic enzymes and superoxide dismutase 2 (SOD2), enhancing the organelle's capacity to neutralize reactive oxygen species. At the same time, cellular NAD+ levels directly regulate genomic integrity through PARP-1. When genotoxic stress or oxidative damage induces single- or double-strand DNA breaks, PARP-1 binds to the damaged termini using its zinc-finger domains. Bound PARP-1 hydrolyzes NAD+ to synthesize extensive, negatively charged poly(ADP-ribose) polymers on itself and adjacent histones. This modification relaxes chromatin architecture and establishes an electrostatic scaffold that recruits base excision repair and homologous recombination complexes. Recent discoveries demonstrate close crosstalk between sirtuins and PARP-1 during DNA repair. PARP-1 recruits SIRT1 to double-strand breaks, where SIRT1 deacetylates the chromatin-remodeling ATPase BRG1 to displace nucleosomes and facilitate homologous recombination. However, because PARP-1 and SIRT1 draw from the same intracellular NAD+ pool, severe NAD+ depletion forces a biological compromise: PARP-1 consumes the scarce remaining cofactor to address DNA damage, leaving sirtuins deactivated. Restoring NAD+ via NR prevents this deficit, enabling concurrent genomic repair and sirtuin-mediated metabolic defense. Quercetin: A Powerful Senolytic and mTOR RegulatorClearing Senescent "Zombie" Cells from TissuesThe accumulation of senescent cells has driven interest in senolytics: molecules that selectively eliminate senescent cells while sparing healthy, non-senescent populations. Senolytic agents exploit a specific vulnerability in senescent cells. Because senescent cells produce cytotoxic, pro-inflammatory SASP factors that would normally induce their own death, they become dependent on upregulated Senescent Cell Anti-Apoptotic Pathways (SCAPs) to survive. The SCAP network involves anti-apoptotic proteins (such as BCL-2 and BCL-xL), the PI3K/Akt kinase cascade, and cyclin-dependent kinase inhibitors.Quercetin is a polyphenolic flavonoid found in capers, red onions, apples, and the flower buds of Sophora japonica. Beyond its classical antioxidant properties, quercetin functions as a senolytic compound that exerts multi-target inhibitory effects across the SCAP network. By inhibiting the upstream PI3K/Akt survival axis and downregulating anti-apoptotic defenses, quercetin disrupts the signaling that protects senescent cells from intrinsic apoptosis. Deprived of these survival signals, senescent cells undergo programmed cell death. Preclinical studies demonstrate that senolytic protocols utilizing quercetin - often combined with the tyrosine kinase inhibitor dasatinib - reduce senescent cell burden across multiple tissues. This targeted clearance lowers circulating SASP factors, attenuates tissue fibrosis, restores endothelial reactivity, and improves functional health span. By removing senescent cells, quercetin mitigates the primary driver of chronic, low-grade inflammaging. Modulating the mTOR Pathway for Optimal AutophagyThe mechanistic Target of Rapamycin (mTOR) is an evolutionarily conserved serine/threonine protein kinase that coordinates cellular metabolism by balancing anabolic growth with catabolic recycling. Operating within two multiprotein complexes - mTORC1 and mTORC2 - the mTOR pathway integrates signals from amino acids, growth factors, and intracellular energy levels. In nutrient-rich environments, mTORC1 promotes protein synthesis, lipogenesis, and cellular growth, while suppressing catabolic breakdown. Conversely, nutrient scarcity downregulates mTORC1, activating autophagy.Autophagy is an intracellular degradation system that packages damaged organelles, misfolded protein aggregates, and biological debris into double-membraned autophagosomes for lysosomal degradation and recycling. A specialized branch of this pathway, mitophagy, selectively targets and clears damaged mitochondria. In modern metabolic conditions characterized by continuous caloric intake, mTORC1 can remain persistently active. This persistent signaling suppresses autophagy, causing damaged organelles and toxic aggregates to accumulate within tissues. Quercetin functions as a natural modulator of mTOR signaling. By inhibiting upstream PI3K/Akt signaling and activating intracellular energy sensors, quercetin attenuates overactive mTORC1, mimicking the metabolic effects of caloric restriction. This down-regulation relieves inhibition on the ULK1 autophagy initiation complex, stimulating both general autophagy and mitophagy. As autophagy proceeds, cells clear protein aggregates and eliminate damaged mitochondria, supporting cellular longevity and proteostasis. Enhancing Absorption: Phytosomes and Dietary FatsDespite the biological activities of quercetin identified in experimental models, its clinical translation has historically been limited by poor oral bioavailability. Raw quercetin aglycone is a crystalline, hydrophobic polyphenol with poor solubility in water and gastrointestinal fluids. When ingested in unformulated powder forms, quercetin molecules aggregate in the gut lumen, resisting dissolution and passive absorption. Consequently, the vast majority of an unformulated dose passes into the colon unabsorbed, where it undergoes microbial degradation without reaching meaningful systemic concentrations.To address these pharmacokinetic limitations, advanced delivery systems such as phytosomes were engineered. A phytosome is a 100% food-grade molecular complex where individual polyphenolic molecules are bound to dietary phospholipids, typically sunflower-derived phosphatidylcholine. Unlike a classical liposome - which encapsulates water-soluble compounds inside an aqueous core enclosed by a lipid bilayer - a phytosome forms an amphiphilic complex at the molecular level. The polar head of the phosphatidylcholine molecule forms hydrogen bonds with the hydroxyl groups of the quercetin molecule, while its lipophilic fatty acid tails extend outward. This structural arrangement shields the polar regions of the flavonoid, creating a lipid-compatible complex that integrates smoothly into the intestinal mucosa.
The Importance of Methylation in Healthy AgingVitamin B-Complex and Choline as Essential Methyl DonorsMethylation is an essential biochemical process occurring billions of times each second across all human tissues. It involves the transfer of a single-carbon unit - a methyl group consisting of one carbon atom bound to three hydrogen atoms - (CH3) - from a donor molecule to diverse recipients, including DNA, RNA, structural proteins, neurotransmitters, and membrane phospholipids. This transfer of one-carbon units is coordinated by the methionine-homocysteine cycle, which sustains genetic stability, detoxification pathways, and cellular repair.At the center of this pathway sits S-adenosylmethionine (SAM), the universal methyl donor in human biology. When a methyltransferase enzyme transfers a methyl group from SAM to an acceptor molecule, SAM is converted into S-adenosylhomocysteine (SAH). SAH functions as a potent competitive inhibitor of intracellular methyltransferases. To maintain functional methylation, SAH is rapidly hydrolyzed into homocysteine, a sulfur-containing amino acid that must be remethylated or cleared through transsulfuration. Homocysteine clearance proceeds through two distinct remethylation pathways. The primary route operates across most tissues via the enzyme methionine synthase, which requires vitamin B12 in its active methylcobalamin form. Methionine synthase transfers a methyl group from 5-methyltetrahydrofolate (5-MTHF, the active form of folate) to homocysteine, regenerating methionine. The ongoing production of 5-MTHF depends on the enzyme methylenetetrahydrofolate reductase (MTHFR), which utilizes riboflavin (vitamin B2) as a cofactor. Alternatively, excess homocysteine can be routed into the transsulfuration pathway by vitamin B6 (as pyridoxal-5'-phosphate) to synthesize cystathionine, cysteine, and ultimately the antioxidant glutathione. A secondary remethylation pathway, active predominantly in hepatic and renal tissues, bypasses folate entirely. In this route, dietary choline is oxidized to betaine (trimethylglycine or TMG). The enzyme betaine-homocysteine S-methyltransferase (BHMT) then transfers a methyl group from betaine directly to homocysteine, yielding methionine and dimethylglycine. When dietary intake of active B-vitamins or choline is insufficient, or when genetic variations like MTHFR polymorphisms reduce pathway flux, the methylation cycle slows. Homocysteine accumulates in circulation, promoting vascular and neurological inflammation, while SAM reserves decline, restricting cellular methylation capacity. Understanding DNA Methylation and Epigenetic HealthEvery somatic cell in an organism carries an identical genetic code. Cellular differentiation and tissue-specific functions are governed by the epigenome: a regulatory layer of chemical modifications that dictates gene expression without altering underlying DNA sequences. DNA methylation represents the primary and most stable epigenetic modification. In this process, DNA methyltransferase (DNMT) enzymes utilize methyl groups donated by SAM to add a methyl tag to cytosine bases adjacent to guanine residues, forming 5-methylcytosine within CpG dinucleotide sites.Under physiological conditions, DNA methylation maintains genomic stability and coordinates transcription. Methylation of promoter regions condenses chromatin, repressing transposable elements and silencing genes inappropriate for a given cell type. Conversely, hypomethylated promoters maintain an open chromatin state, allowing transcription factors to bind and initiate gene expression. During biological aging, this epigenetic landscape undergoes progressive dysregulation, a phenomenon termed "epigenetic drift". Aging cells experience global hypomethylation alongside focal hypermethylation of specific gene promoters. Global loss of methyl tags destabilizes the genome, activating retrotransposons and pro-inflammatory pathways. Simultaneously, hypermethylation at targeted promoter sites silences critical tumor suppressor genes and DNA repair complexes. This systematic change in DNA methylation patterns is consistent across populations, allowing researchers to develop molecular "epigenetic clocks". Algorithms such as the Horvath clock, PhenoAge, and GrimAge quantify biological age by profiling the methylation status of specific CpG sites across the genome. These clocks assess whether individuals are aging faster or slower than their chronological years. Ensuring a steady supply of methyl donors and preventing unnecessary SAM depletion supports DNMT activity, maintaining epigenetic patterns and genomic stability. How the Methylation Cycle Impacts Energy and Cognitive FocusBeyond long-term epigenetic regulation, the methylation cycle directly modulates immediate biochemical processes that govern daily energy, neurotransmission, and cognitive focus. Compromised methylation capacity frequently manifests as cognitive slowing, executive fatigue, and reduced physical stamina.A major consumer of methyl reserves is the endogenous synthesis of creatine. Approximately 40% of all SAM-derived methyl groups in the human body are utilized by guanidinoacetate N-methyltransferase (GAMT) in the liver to synthesize creatine. Creatine then translocates to the brain and skeletal muscle, where it is phosphorylated into phosphocreatine. Phosphocreatine functions as a rapid energy buffer, donating a high-energy phosphate group to regenerate ADP into ATP in milliseconds during demanding physical or cognitive tasks. When methyl donor availability falls, endogenous creatine synthesis drops, depleting phosphocreatine reserves and increasing susceptibility to neuromuscular and cognitive fatigue. Methylation is equally central to central nervous system architecture. SAM provides methyl groups to convert phosphatidylethanolamine into phosphatidylcholine, the predominant phospholipid comprising neuronal cell membranes and the myelin sheaths that insulate axons. Intact myelin preserves rapid action potential conduction throughout the nervous system. Furthermore, free choline derived from this pathway is the direct precursor to acetylcholine, the neurotransmitter required for attention, working memory, and learning. The methylation cycle also governs monoamine neurotransmitter metabolism. SAM is required for the synthesis of adrenaline (epinephrine) from noradrenaline, while catechol-O-methyltransferase (COMT) relies on SAM to degrade dopamine and norepinephrine within the prefrontal cortex. Sluggish methylation disrupts this balance, contributing to cognitive fatigue, mood variability, and impaired mental performance. Building a Comprehensive Longevity ProtocolSynergizing NR, Quercetin, and Methylated B-VitaminsLongevity supplementation often falters when single molecules are administered in isolation, ignoring interconnected metabolic pathways. Designing an effective cellular longevity protocol requires combining complementary mechanisms that reinforce one another while preventing secondary metabolic deficits. The combination of Nicotinamide Riboside, Quercetin Phytosome, and Methylated B-Vitamins illustrates this multi-target synergy.This synergy is grounded in the direct biochemical intersection between the NAD+ salvage pathway and the methylation cycle. When high-dose NR is supplemented to boost systemic NAD+, sirtuins and PARP enzymes consume the newly synthesized cofactor, generating substantial quantities of free nicotinamide (NAM). This intracellular nicotinamide faces two primary metabolic fates: it can be recycled back into NAD+ through the NAMPT-dependent salvage loop, or it can be cleared via methylation. When the influx of nicotinamide exceeds salvage recycling capacity, the excess is cleared to avoid feedback inhibition of sirtuin enzymes. To accomplish this, the enzyme nicotinamide N-methyltransferase (NNMT) transfers a methyl group from SAM directly onto nicotinamide, forming 1-methylnicotinamide (1-MNA/MNAM), which is subsequently excreted in urine. Prolonged, high-dose precursor administration without nutritional methyl support can elevate NNMT flux, depleting intracellular SAM reserves. As methyl groups are consumed clearing nicotinamide, the cellular SAM-to-SAH ratio falls, which can elevate circulating homocysteine and reduce methyl availability for DNA methylation and neurotransmitter synthesis. Co-administering a fully methylated B-complex alongside choline or betaine addresses this potential bottleneck. Providing active methyl donors (such as 5-MTHF, methylcobalamin, and betaine) maintains the one-carbon donor pool. Even during increased NNMT activity, SAM pools remain stable, protecting DNA methylation fidelity and maintaining homocysteine within safe parameters. Quercetin reinforces this protocol through complementary mechanisms. By clearing senescent cells and reducing SASP-mediated inflammation, quercetin downregulates CD38, the primary enzyme responsible for age-related NAD+ degradation. Suppressing CD38 prevents unnecessary breakdown of newly synthesized NAD+, enhancing the efficiency of NR supplementation. Furthermore, while NR provides the NAD+ necessary to activate SIRT1-driven mitochondrial biogenesis, quercetin concurrently modulates mTORC1 to stimulate autophagy. This coordinated action ensures that newly generated mitochondria operate in an environment cleared of proteotoxic cellular debris. The Crucial Role of Magnesium Glycinate and Zinc in Cellular FunctionLongevity protocols require essential mineral cofactors to function efficiently. Without adequate divalent minerals acting as enzymatic cofactors and structural stabilizers, metabolic longevity pathways cannot operate at full capacity. Among these, magnesium and zinc are required for cellular repair, genomic stability, and energy production.Magnesium serves as an obligatory cofactor in over 300 enzymatic reactions, primarily through its interaction with ATP. In biological systems, ATP exists predominantly as a chelate with a divalent magnesium ion, forming biologically active Mg2+ -ATP. Every enzymatic reaction that synthesizes, transfers, or consumes cellular energy - including the enzymes of the NAD+ salvage pathway (NRK and NMNAT) and DNA polymerases - strictly requires Mg2+ -ATP as its substrate. Magnesium deficiency impairs these phosphorylation reactions, reducing the cellular utilization of NAD+ precursors. Additionally, magnesium is an essential cofactor for the enzymes that activate dietary B-vitamins into their active forms. Supplying magnesium as magnesium glycinate provides high gastrointestinal bioavailability, minimal laxative effect, and yields glycine to support inhibitory neurotransmission and restful sleep. Zinc serves as a vital structural component for more than 3,000 human transcription factors and enzymatic proteins. Its most prominent structural role in longevity occurs within zinc-finger motifs. These are specialized protein conformations stabilized by a zinc ion coordinated to cysteine and histidine residues. The DNA damage sensor PARP-1 utilizes three zinc-finger domains to identify, track, and physically bind to single- and double-strand DNA breaks. Without adequate intracellular zinc, PARP-1 cannot properly assemble or dock onto damaged chromosomes, impairing DNA repair and increasing genomic instability. Zinc is also an obligatory structural component of copper/zinc superoxide dismutase (Cu/Zn-SOD or SOD1), the primary cytosolic antioxidant enzyme that dismutates superoxide radicals into hydrogen peroxide, protecting mitochondrial membranes and nuclear DNA from premature senescence. Integrating Prebiotics (like Acacia and Inulin) for Gut-Derived Longevity MarkersA comprehensive cellular longevity framework must extend beyond somatic tissues to encompass the gut microbiome. The intestinal microbiome functions as a central regulator of systemic inflammatory tone, immune development, and metabolic signaling. Age-associated dysbiosis - characterized by the loss of beneficial commensals and an overgrowth of pathobionts - frequently leads to breakdown of the intestinal barrier.The gut epithelium consists of a single-cell monolayer sealed by tight junction proteins, including zonula occludens-1 (ZO-1), occludin, and claudins. When this physical barrier is disrupted by poor dietary fiber intake or dysbiosis, gut permeability increases. This allows lipopolysaccharide (LPS), a component of the outer membrane of Gram-negative bacteria, to enter the portal and systemic circulation. The resulting "metabolic endotoxemia" activates Toll-like receptor 4 (TLR4) on immune cells, inducing NF-kB and systemic pro-inflammatory cytokine production. This persistent gut-derived inflammation exacerbates the SASP, accelerates tissue senescence, upregulates CD38, and drains systemic NAD+ reserves.
These short-chain fatty acids, particularly butyrate, exert direct protective effects on systemic longevity. Butyrate provides the primary metabolic fuel for colonic epithelial cells, supplying more than 70% of their baseline energy needs and supporting mitochondrial function within colonocytes. Furthermore, SCFAs upregulate the expression of epithelial tight junction proteins (ZO-1, occludin, and claudin-1), restoring intestinal barrier integrity and preventing the translocation of inflammatory LPS into systemic circulation. Systemically absorbed butyrate also functions as an endogenous histone deacetylase (HDAC) inhibitor, suppressing pro-inflammatory gene expression and supporting regulatory T cell (T_reg) development. Reducing metabolic endotoxemia dampens systemic inflammation, protecting vascular function and preventing premature NAD+ depletion. Conclusion: The Integrated Cellular Longevity MatrixCellular longevity is achieved not by addressing isolated biomarkers in isolation, but by systematically supporting interconnected biological pathways. As bioenergetic capacity declines, cellular senescence accelerates, epigenetic patterns degrade, and gut barrier integrity weakens. A comprehensive approach addresses these biological vulnerabilities simultaneously.
Quercetin Phytosome clears senescent cells and modulates mTORC1, stimulating autophagy while dampening the inflammatory SASP cascade that accelerates CD38-mediated NAD+ destruction. Methylated B-vitamins, active folate, and choline replenish SAM reserves, balancing the methyl requirements of NNMT-mediated nicotinamide clearance, preserving epigenetic DNA methylation, and maintaining neurotransmitter production. Magnesium glycinate and zinc provide the structural and catalytic foundation required for ATP utilization, B-vitamin activation, and PARP-1 zinc-finger DNA repair docking. Finally, prebiotic fibers generate short-chain fatty acids like butyrate, reinforcing the intestinal barrier and preventing metabolic endotoxemia from fueling systemic inflammation. By coordinating energy replenishment, cellular waste clearance, epigenetic maintenance, and the suppression of systemic inflammation, this unified approach directly addresses the underlying drivers of cellular aging to support long-term physiological vitality.
--
(https://vitanetonline.com:443/forums/Index.cfm?CFApp=1&Message_ID=6650) Choline: The Liver’s Fat-Export Engine 
Date:
June 24, 2026 11:48 AM
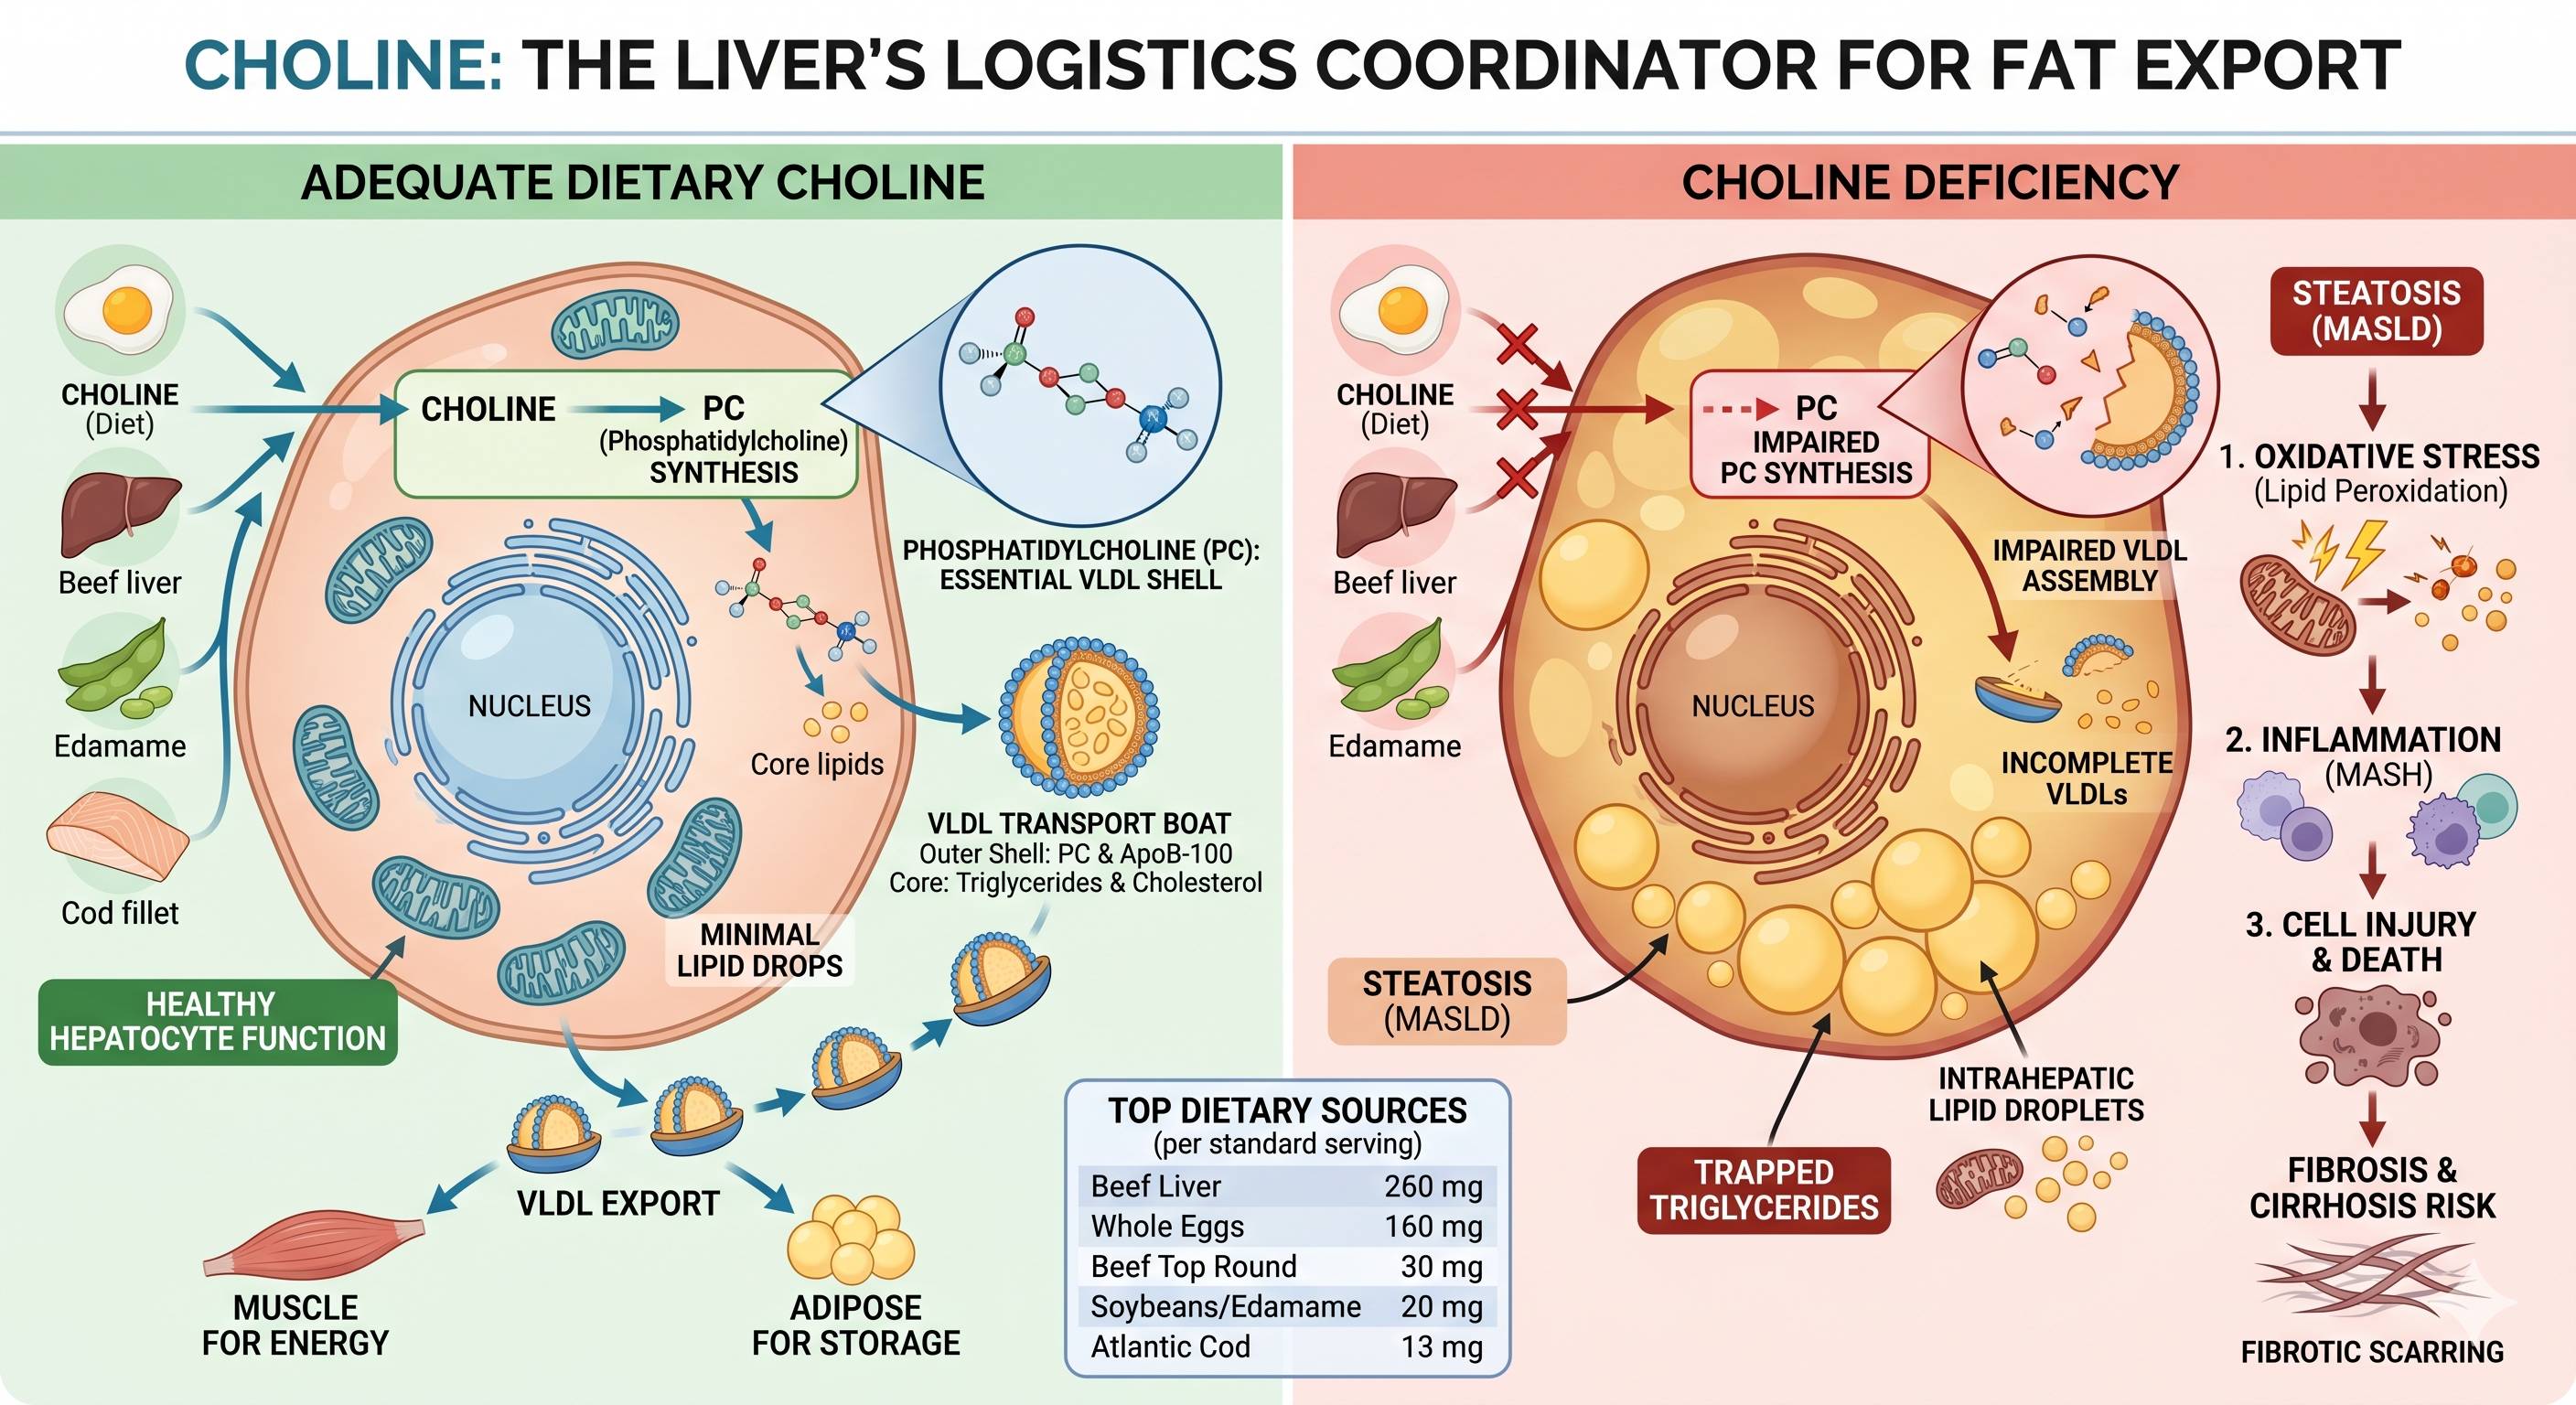
Without sufficient choline, the liver's ability to process and export fats grinds to a halt. The Liver’s Fat-Export MechanismThe primary reason the liver requires choline comes down to a specific molecule: phosphatidylcholine (PC).When you eat fats or liberate stored fatty acids, your liver processes them into triglycerides. To move these triglycerides out of the liver and deliver them to muscle or adipose (fat) tissue for energy or storage, the liver has to package them into transport vehicles called VLDL (Very Low-Density Lipoproteins). Phosphatidylcholine forms the essential outer shell of these VLDL transport "boats." If you don't have enough choline, your liver cannot manufacture this outer shell, meaning the boats cannot be built, and fat cannot leave the liver. What Happens When Choline is Deficient?When dietary choline drops below critical levels, a highly predictable cascade of liver dysfunction occurs:
Dietary Benchmarks & SourcesTo prevent fat accumulation and support proper VLDL export, the Food and Nutrition Board establishes specific Adequate Intake (AI) targets, though optimal metabolic and athletic performance may require a more robust baseline.
Summary: Choline acts as a critical logistics coordinator for liver health, serving as a non-negotiable raw material for exporting fat out of the organ. Specifically, the liver relies on choline to produce phosphatidylcholine, a molecule that forms the essential outer shell of Very Low-Density Lipoproteins (VLDL). Think of VLDLs as transport boats designed to safely carry triglycerides out of the liver and deliver them to the rest of the body for energy or storage. Without an adequate dietary supply of choline, the liver simply cannot build these outer shells, causing the entire transport system to grind to a halt and leaving processed fats stranded inside liver cells. When this export mechanism stalls, the stranded fat quickly triggers a highly predictable cascade of liver dysfunction. This buildup leads directly to fat accumulation - historically known as fatty liver disease and now classified as MASLD - which quickly overwhelms the cellular mitochondria. The resulting oxidative stress and cellular injury spark chronic inflammation (MASH), causing liver cells to die and potentially leading to permanent scarring or fibrosis over time. Furthermore, a severe shortage of dietary choline starves the body's methylation cycle, forcing it to deplete vital SAMe pools in a desperate, structurally taxing attempt to manufacture its own phosphatidylcholine.
(https://vitanetonline.com:443/forums/Index.cfm?CFApp=1&Message_ID=6643) Beyond Choline: How DMAE Flushes Cellular Waste for Longevity
Date:
May 21, 2026 03:43 PM
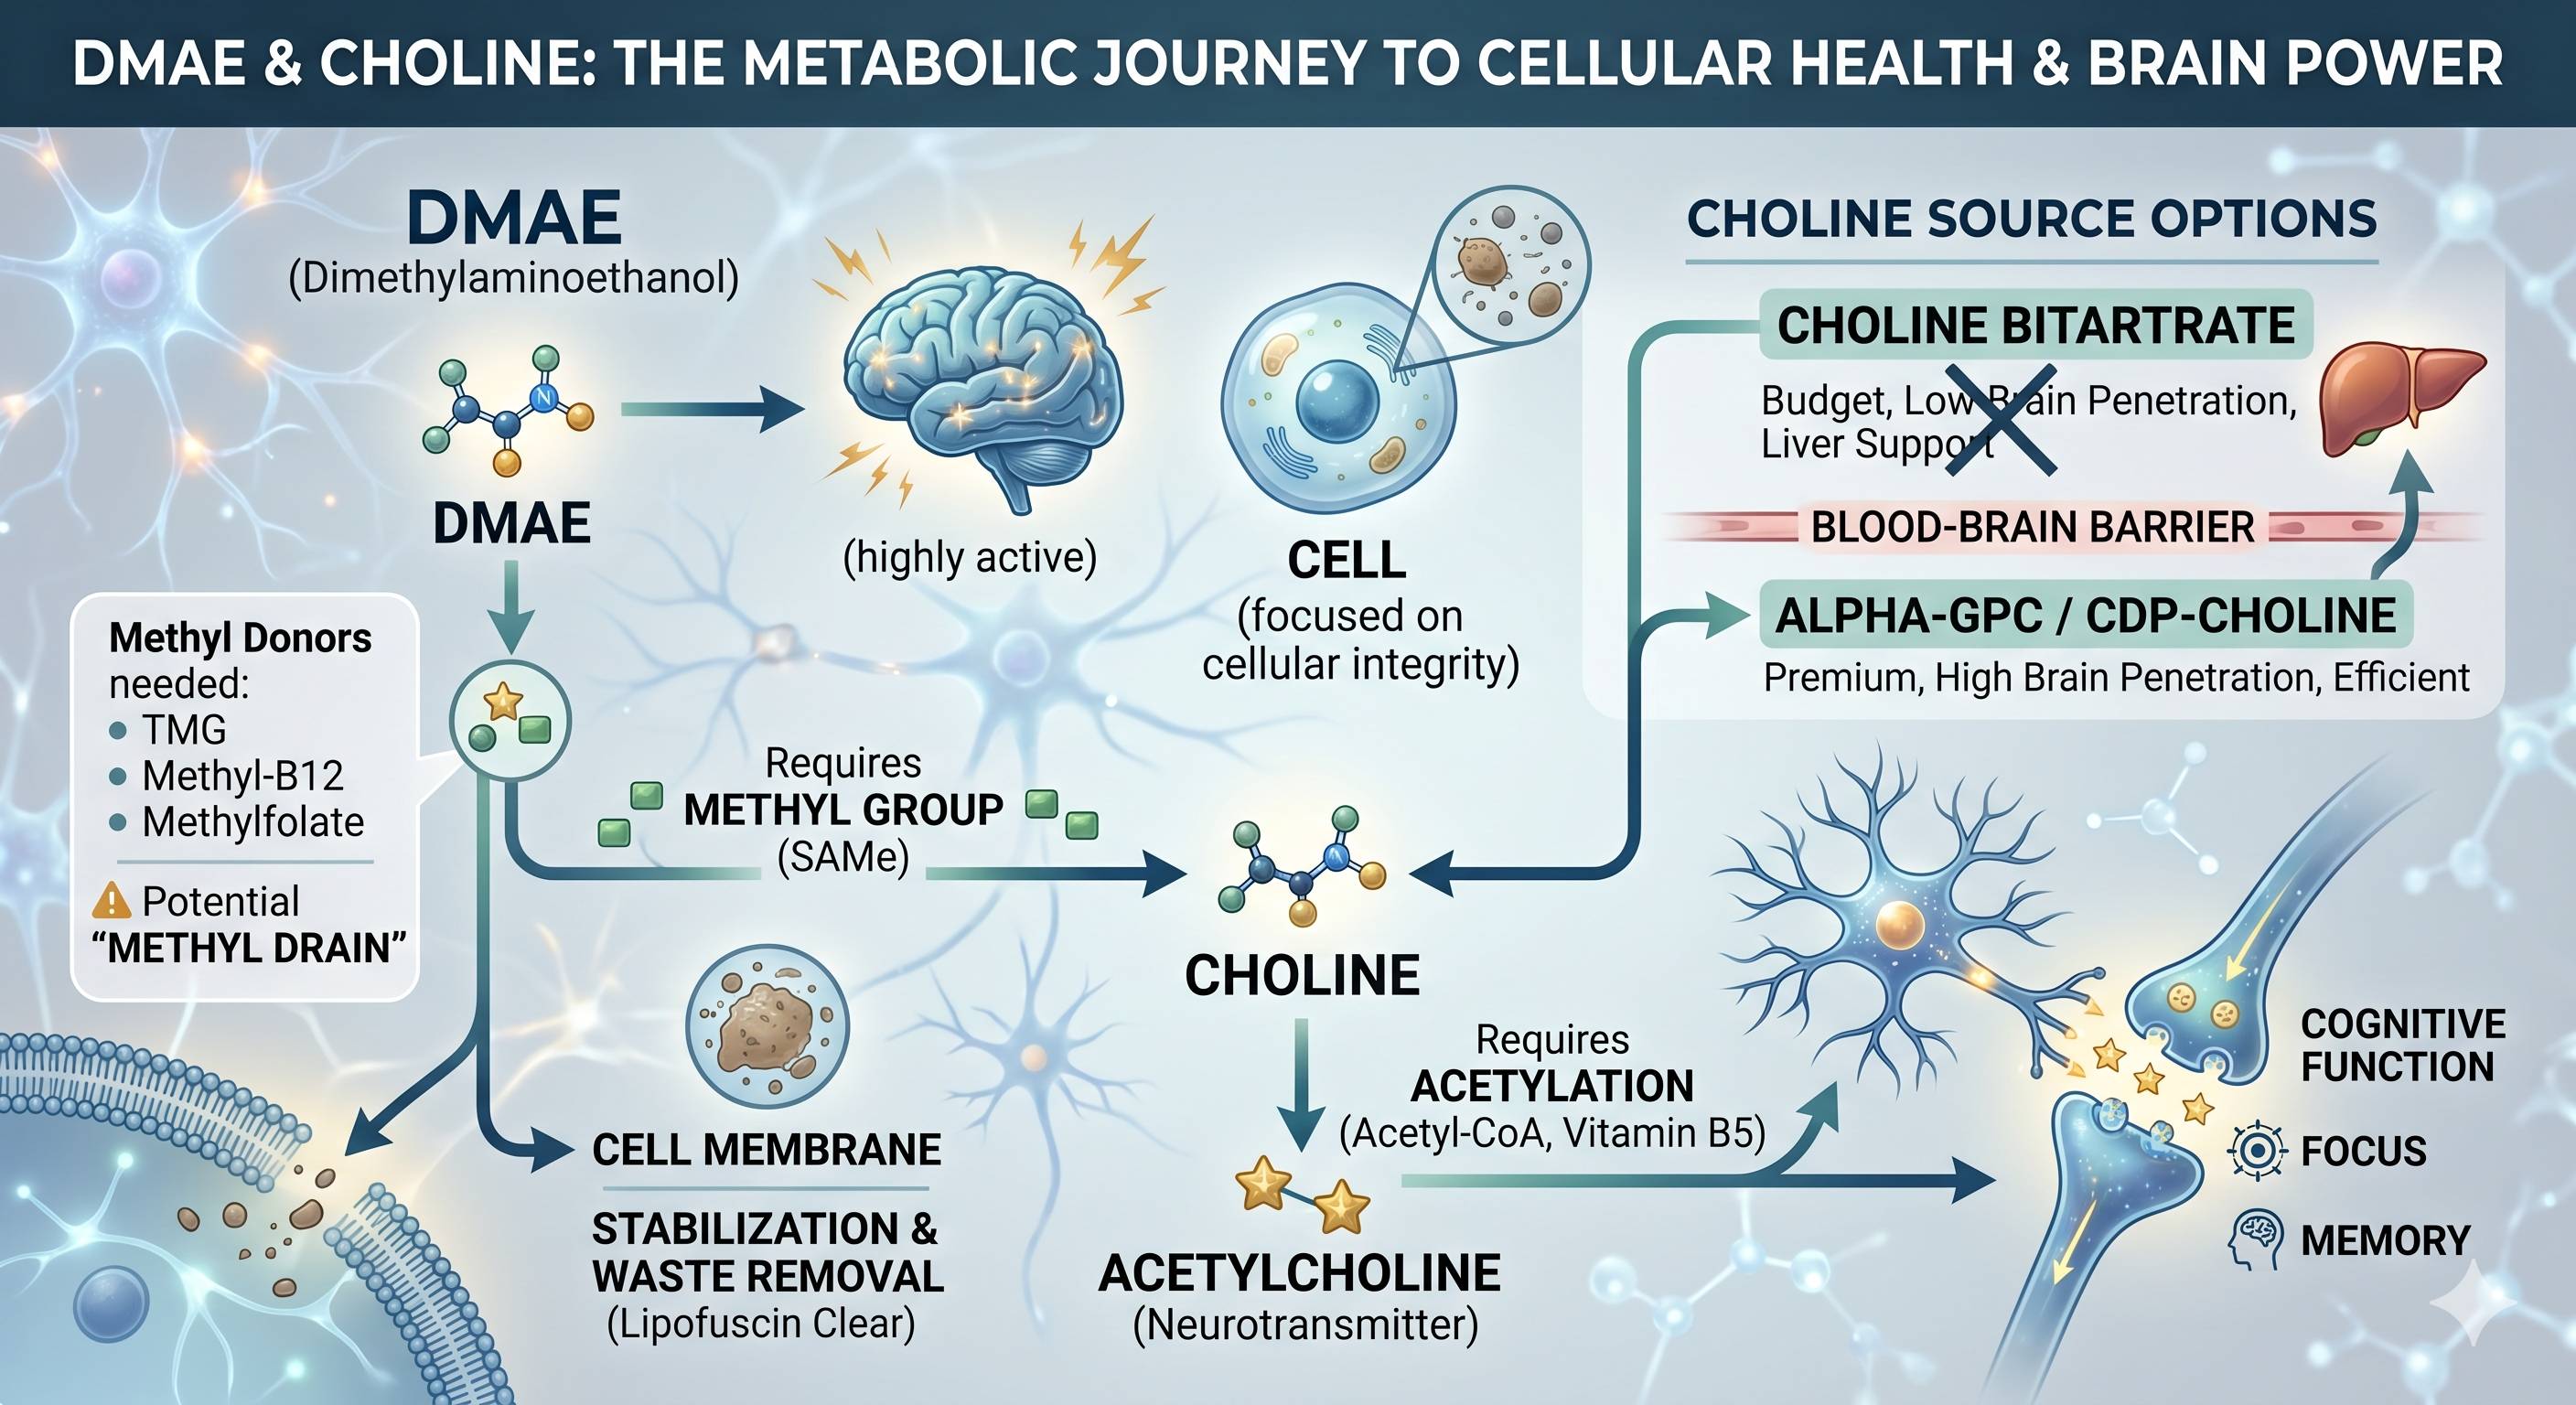
How is DMAE and Methylation connected?Understanding the relationship between DMAE (dimethylaminoethanol) and the methylation cycle is a nuanced dive into nutritional biochemistry. While DMAE is a popular compound for cognitive and cellular health, its interaction with the body’s methyl pool is a critical factor to consider.DMAE and the Methylation ConnectionBiochemically, it is often considered important to support the methylation cycle when supplementing with DMAE. This is due to how the body processes DMAE to create choline.The "Methyl Drain"DMAE is a precursor to choline, but the conversion process requires the addition of methyl groups. Specifically, the body uses SAMe (S-adenosylmethionine), the universal methyl donor, to methylate DMAE into choline.If you supplement with high doses of DMAE without adequate methyl support, you may inadvertently "drain" your methyl pool. This can lead to:
Strategic Co-factorsTo balance this, many researchers suggest pairing DMAE with methyl donors to ensure the cycle remains fluid. Key co-factors include:
How DMAE Supports LongevityDMAE is often categorized as a "longevity" nutrient because of its impact on cellular waste and membrane integrity.1. Reduction of LipofuscinOne of the most cited longevity benefits of DMAE is its ability to reduce lipofuscin (often called "age spots" or "wear-and-tear pigment"). Lipofuscin is a metabolic waste product that accumulates in the heart, brain, and skin cells over time. High levels of lipofuscin can impair cellular function; DMAE helps the body flush these deposits, potentially slowing the cellular aging process.2. Membrane StabilizationDMAE acts as a free radical scavenger specifically within the cell membranes. By protecting the lipid bilayer from oxidative stress, it helps maintain membrane fluidity. Flexible, healthy membranes are essential for efficient nutrient transport into the cell and waste removal out of the cell.3. Acetylcholine PrecursorWhile its primary longevity mechanism is cellular, DMAE’s role as a precursor to acetylcholine supports cognitive longevity. Maintaining healthy levels of this neurotransmitter is vital for focus, memory, and muscle control as the nervous system ages.4. Skin Health and "Lifting"In the context of physical appearance and anti-aging, DMAE is known for its "firming" effect. It is thought to increase the tone of the underlying skin muscles, providing a subtle "lift" and reducing the appearance of fine lines and sagging.Summary: While DMAE offers potent cellular cleaning and membrane protection, it functions best when the body has a robust supply of methyl donors to handle its conversion without taxing the system. Does DMAE efficiently convert to Acetylcholine?The conversion of DMAE (dimethylaminoethanol) into the neurotransmitter acetylcholine is a multi-step biochemical process. Because DMAE is not a direct precursor to acetylcholine, it must first be integrated into the phospholipid metabolism of the cell.To make this conversion efficient, the body requires specific methyl donors and enzymatic co-factors. The Two-Step Conversion Process1. The Methylation Step: DMAE to CholineDMAE is essentially a "dimethylethanolamine." To become choline (trimethylethanolamine), it requires the addition of a third methyl group.
2. The Acetylation Step: Choline to AcetylcholineOnce DMAE has been methylated into choline (or integrated into phosphatidylcholine), it must be acetylated by the enzyme Choline Acetyltransferase (ChAT).
Optimal Co-factors for EfficiencyTo maximize the "brain-boosting" effects of DMAE, the following nutrients are typically used to support the pathway:
The "Choline Sparer" EffectIt is worth noting that DMAE is often more efficient at crossing the blood-brain barrier than standard choline. However, because it "competes" for the same transport systems and enzymes, taking it alongside a high-quality choline source and a B-complex ensures that the body isn't forced to choose between building cell membranes and creating neurotransmitters.What is it better to consume choline or DMAE?Deciding between DMAE and choline depends entirely on whether your priority is systemic health (liver and cell membranes) or cognitive performance (acetylcholine production and focus). While choline is technically a "cheaper" raw material, the "efficiency" changes once you factor in how the body processes each compound.The Efficiency ComparisonEfficiency in this context is usually measured by how well a compound crosses the blood-brain barrier (BBB) and how many metabolic steps are required to reach the final goal: Acetylcholine.1. Choline Bitartrate (The "Budget" Option)
2. DMAE (The "Specialist")
3. Alpha-GPC & CDP-Choline (The "Premium" Options)
Cost-Efficiency Analysis
Which is better?
Summary: Supplementing with DMAE (dimethylaminoethanol) creates a unique biochemical dynamic in the body, particularly regarding cognitive performance and cellular longevity. While DMAE crosses the blood-brain barrier effectively, its conversion into the crucial neurotransmitter acetylcholine requires an additional methyl group, a process that can significantly drain the body's universal methyl donor, SAMe. To prevent this "methyl drain" and avoid issues like elevated homocysteine, it is highly beneficial to pair DMAE with strategic co-factors and methyl donors such as TMG (trimethylglycine) and methylated B-vitamins. Beyond its role in acetylcholine production, DMAE supports longevity through distinct cellular mechanisms that standard choline cannot replicate, specifically by neutralizing free radicals to stabilize cell membranes and flushing out lipofuscin, an age-related cellular waste product. When evaluating whether to replace DMAE with direct choline supplementation for cost and efficiency, the decision hinges on your specific physiological goals. Standard budget options like choline bitartrate are highly cost-effective for general liver health and systemic cell structures, but they suffer from very poor brain penetration. Conversely, premium forms like Alpha-GPC and CDP-Choline are far more efficient for cognitive optimization and memory because they cross the blood-brain barrier effortlessly without taxing the body's methylation cycle. Ultimately, while cheap choline salts cannot replicate DMAE's unique cellular cleaning properties, choosing a premium choline source is a more direct and biochemically efficient path for raw cognitive enhancement, especially if you want to avoid the multi-step metabolic cost associated with converting DMAE.
(https://vitanetonline.com:443/forums/Index.cfm?CFApp=1&Message_ID=6638) Is NAD+ the "Missing Link" in Modern Longevity Science?
Date:
April 11, 2026 12:24 PM
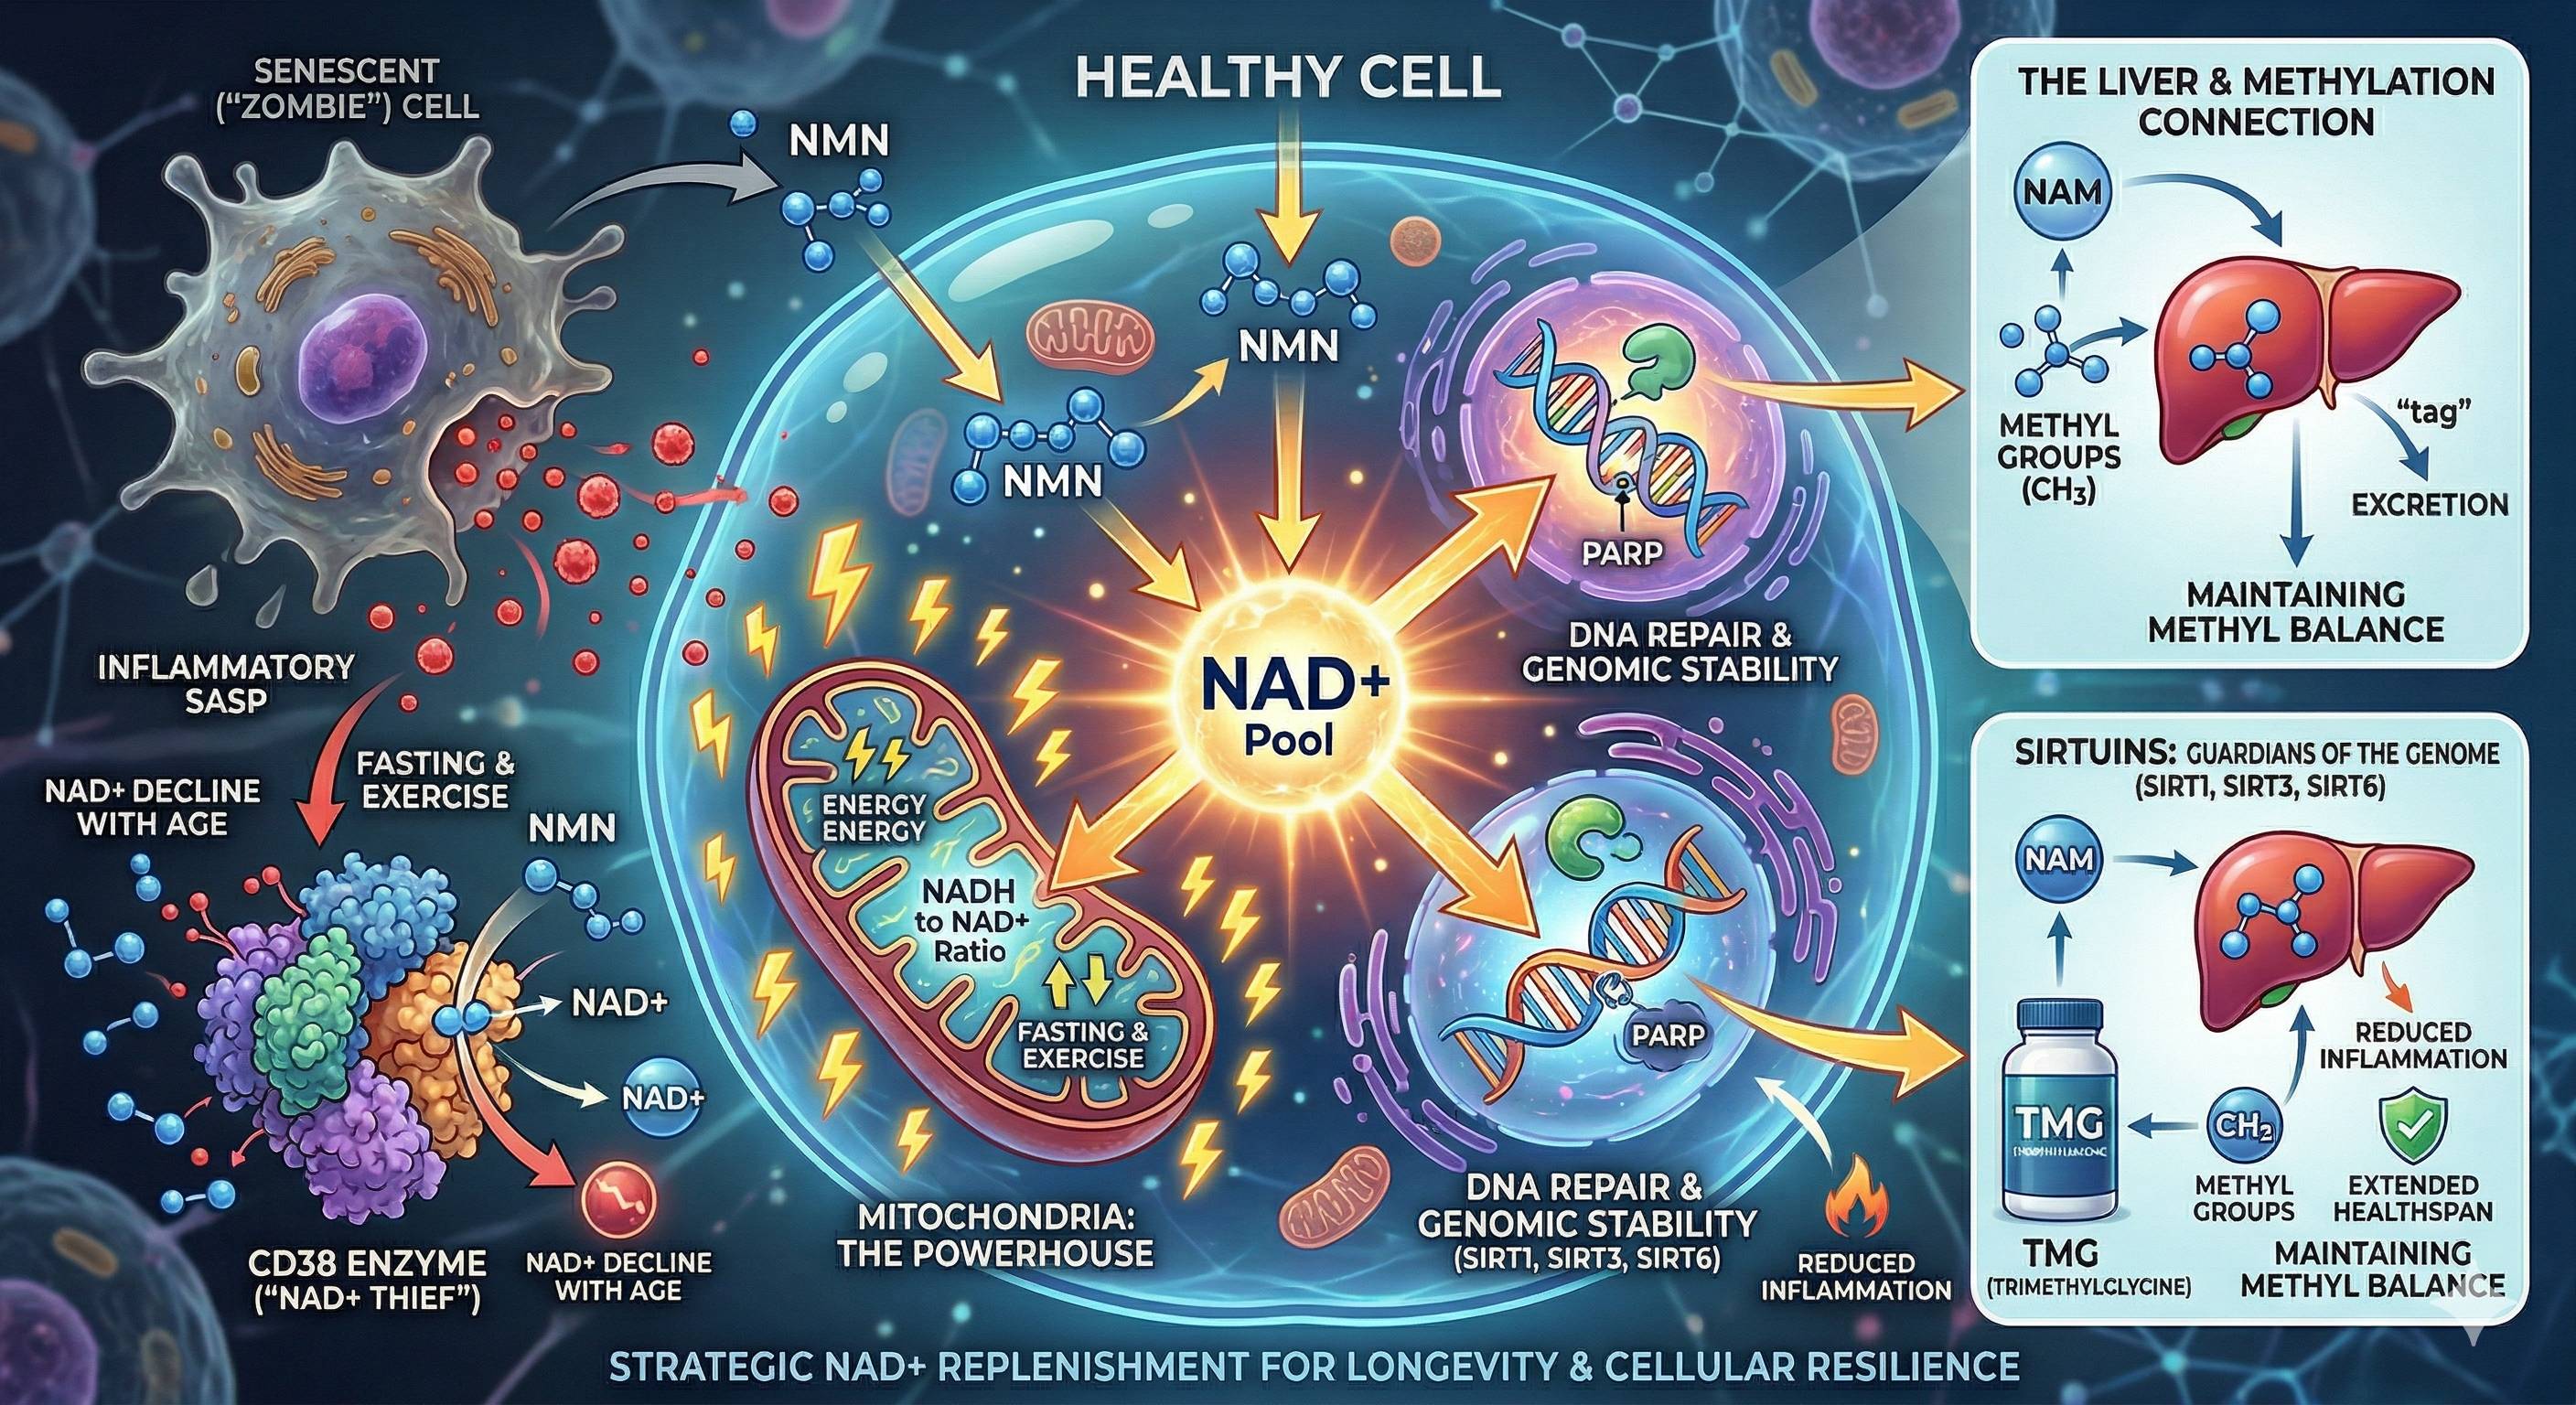
How NMN Boosts NAD+ LevelsNMN is essentially a "building block" for NAD+. The process by which the body converts NMN into NAD+ is relatively direct, making it an efficient supplement for raising levels.
Why Boost NAD+? The Biological BenefitsNAD+ is a "coenzyme" found in every single living cell. It is required for over 500 different enzymatic reactions. Here are the primary reasons why maintaining high NAD+ levels is critical for health:1. Cellular Energy Production (ATP)NAD+ plays a starring role in the mitochondria, the powerhouses of the cell. It acts as an electron carrier in the process of turning nutrients from food into ATP (Adenosine Triphosphate), which is the primary energy currency of the body. Without enough NAD+, cellular energy production falters, leading to fatigue and decreased metabolic function.2. DNA Repair and StabilityOur DNA is constantly under attack from UV radiation, pollutants, and metabolic byproducts. To fix this, the body uses enzymes called PARPs (Poly ADP-Ribose Polymerases). These enzymes are strictly "NAD+-dependent," meaning they cannot function without consuming NAD+. By boosting NAD+, you provide the fuel necessary for cells to repair genetic damage effectively.3. Activation of Sirtuins (The "Longevity Genes")Sirtuins are a family of proteins responsible for cellular health, aging, and DNA protection. They are often called "guardians of the genome."
4. Metabolic Health and Insulin SensitivityResearch indicates that increasing NAD+ through NMN can improve glucose metabolism and insulin sensitivity. This is particularly relevant for maintaining a healthy weight and reducing the risk of age-related metabolic disorders.5. Circadian Rhythm RegulationNAD+ levels fluctuate throughout the day and help regulate the body’s internal clock (the circadian rhythm). Maintaining healthy levels helps ensure that your cells know when to be active and when to undergo repair, which can improve sleep quality and daytime alertness.Note: While NMN is widely studied in animal models with promising results regarding lifespan and healthspan, human clinical trials are ongoing to fully confirm long-term efficacy and optimal dosage. Always consult a healthcare professional before starting new supplements. Furthermore:While the basics of NMN and NAD+ focus on energy and aging, there is a much deeper layer to this "master molecule" that involves cellular theft, "zombie" cells, and the critical importance of a process called methylation.Here are the vital pieces of the NAD+ puzzle that aren't usually mentioned: 1. The "NAD+ Thieves": CD38 and InflammationBoosting NAD+ with NMN is only half the battle; the other half is stopping the "thieves" that steal it.
2. The Methylation Connection (Why you need TMG)This is perhaps the most important practical detail for anyone taking NMN.
3. The "Double-Edged Sword" of CancerNAD+ is essential for all living cells, including cancer cells.
4. Regulatory Status (The 2026 Update)The legal status of NMN has been a rollercoaster. In late 2022, the FDA initially ruled that NMN could not be marketed as a supplement because it was being investigated as a drug. However, as of late 2025, the FDA issued updated guidance essentially reinstating NMN’s status as a legal dietary ingredient, clearing the way for its continued sale in the U.S.5. Natural Boosters and "Sirtuin Activators"Supplementing NMN is just one way to boost the system. You can also:
Are you planning to focus your blog post on the anti-aging benefits for a general audience, or would you like to lean more into the technical "biohacking" side of science?
(https://vitanetonline.com:443/forums/Index.cfm?CFApp=1&Message_ID=6636) Betaine aka TMGs Many Benefits as a Methyl Donor
Date:
October 21, 2022 12:29 PM
Methylation is a process in the body that helps to produce DNA, keep hormones in balance, and support detoxification. You need methyl groups for this process to happen, and TMG (trimethylglycine) donates methyl groups to help with methylation. TMG also supports cardiovascular health and cognitive function while reducing inflammation. Not sure if TMG is right for you? Keep reading to learn more about this methyl donor’s benefits. Benefit #1: TMG Promotes Healthy Methylation As we just touched on, one of the main benefits of TMG is that it promotes healthy methylation. As we know, methylation is important for DNA production, hormone balance, and detoxification support. TMG provides the extra push of methyl groups needed to keep these processes running smoothly which is why it’s often taken as a supplement. If you have difficulty methylating or you have a genetic mutation that affects methylation, TMG may be a good supplement for you to consider. Check with your doctor first, of course! Benefit #2: TMG Supports Cardiovascular Health TMG also known as betaine, can improve cardiovascular health by reducing homocysteine levels. Homocysteine is an amino acid that at high levels has been linked to heart disease. While more research needs to be done in this area, initial studies are promising. In one study of nearly 500 participants, researchers found that those who took TMG had lower levels of homocysteine than those who didn’t supplement with the nutrient [1]. While this doesn’t prove causation, it’s an interesting correlation worth noting—especially given that other studies have found similar results [2]. benefit #3: May Reduce Inflammation Chronic inflammation has been linked to many diseases such as heart disease, arthritis, and even cancer. Some studies suggest that TMG may help reduce inflammation levels in the body [3]. In one animal study, rats given TMG had reduced lung inflammation compared to rats who weren’t given the supplement [4]. More research needs to be done in this area but initial studies are promising. In Summary: TMG is a nutrient known as a methyl donor which means it provides methyl groups needed for processes like DNA production and hormone balance. It also supports cardiovascular health and cognitive function while reducing inflammation. If you have difficulty methylating or you have a genetic mutation that affects methylation, TMG may be a good supplement for you to consider. Check with your doctor first!
(https://vitanetonline.com:443/forums/Index.cfm?CFApp=1&Message_ID=6514) Few reason that tell you why you should consume TMG regularly
Date:
January 11, 2014 05:20 PM
TMG or Trimethylglycine
Benefits of TMG If we talk about benefits of TMG or Trimethylglycine it is responsible for proper liver functioning, body detoxification, cellular replication, carinitine manufacturing, protection from diabetes and protection of kidney from any kind of damage. Other than this it can also help you to avoid a lot of genetic problems by regulating the genetic transcription process that is responsible for the creation of all the chemicals in body. In addition to this many researchers also believe it that TMG can improve the autonomous nervous system of body and if it is unbalanced, then it can help you to restore the system. Since TMG is responsible for cell duplication so if someone is facing problem of cell synthesis TMG can help those people as well in restoration of their regular body structure. In short we can say that TMG is one of the most important substances of human body that is very helpful for proper functioning of body in a very effective manner. Recommendation Therefore, it is recommended that if you are lacking for TMG you should take it as a dietary supplement so you can increase its count in your body and you can avoid these health problems and you can restore proper functioning of your body. However, it is also very important that you should take the TMG according to specific guideline suggested by your doctor and if you ever find any problem while consuming it, make sure you communicate your doctor about those problems.
(https://vitanetonline.com:443/forums/Index.cfm?CFApp=1&Message_ID=2961) Supports Healthy Blood Pressure
Date:
April 18, 2008 03:59 PM
3 tablets contain: Suggested Use: 3 tablets mid-morning and 2 tabs mid-afternoon between meals on an empty stomach.
(https://vitanetonline.com:443/forums/Index.cfm?CFApp=1&Message_ID=1763) TMG Fact Sheet
Date:
December 07, 2005 02:13 PM
TMG Fact SheetNeil E. Levin, CCN, DANLA 03/07/05LIKELY USERS: People with high homocysteine levels; People with risks of developing Alzheimer’s Disease; People needing greater metabolism of fats; People with liver detoxification challenges; People consuming alcohol KEY INGREDIENTS: TMG is composed of three methyl groups attached to a glycine atom. It can “donate” methyl groups. MAIN PRODUCT FEATURES: TMG is a metabolite of the B vitamin family product called Choline. Choline has 4 methyl groups, TMG has 3 and DMG has 2. These substances plus Folic acid, Vitamin B-12 and SAM-e are all methyl donors. Methyl donors can contribute methyl groups to biological processes such as liver function, detoxification and cellular replication (production of new cells). Methylation protects the kidneys and stimulates production of the fat-transporting molecule l-carnitine. TMG helps the liver metabolize fats, preventing the accumulation of fats in the liver. It also helps to detoxify chemicals in the liver, while protecting the liver from being damaged by those chemicals. Methylation with TMG helps to convert the dangerous, inflammatory chemical homocysteine into the amino acid methionine. TMG may lower homocysteine when B-6, B-12 and folic acid cannot. ADDITIONAL PRODUCT INFORMATION: TMG is also known as Betaine and is a component of Betaine hydrochloride (Betaine HCl), a stomach acid supplement that is very acidic. But Betaine HCl is not used in the same way as TMG. TMG is not highly acidic and will not supplement low stomach acid. TMG may be useful for autistic children, along with B-6 and magnesium. It may also be useful in strengthening the body’s immune response against pathogenic bacteria. There is very preliminary evidence that TMG and methyl donors may help against some forms of seizures. DMG has been used as a sports supplement. TMG is 50% more effective than DMG in any application where the methyl groups are useful. Otherwise, they can used interchangeably. SERVING SIZE & HOW TO TAKE IT: One serving per day, or up to 6,000 mg., as needed. COMPLEMENTARY PRODUCTS: SAM-e, Milk Thistle (Silymarin), Dr. Verghese’s Liver Detoxifier & Regenerator, Antioxidants, NAC, Homocysteine Regulators, D-Flame, Detox Support CAUTIONS: Pregnant and lactating women and people using prescription drugs should consult their physician before taking any dietary supplement. People with Parkinson’s or taking L-dopa should not use methyl donors like TMG without a physician’s specific approval and supervision. There are no other known drug interactions with TMG. This information is based on my own knowledge and references, and should not be used as diagnosis, prescription or as a specific product claim. This is not an official publication by any company, nor has this information been screened or approved by the FDA or any private company. Disclaimer: These statements have not been evaluated by the FDA. This product is not intended to diagnose, treat, cure or prevent any disease. REFERENCES: General: Craig SA. Betaine in human nutrition. Am J Clin Nutr. 2004 Sep;80(3):539-49. Review. PMID: 15321791 Methylation: Barak AJ, Tuma DJ. Betaine, metabolic by-product or vital methylating agent? Life Sci 1983;32:771-4 [review]. Benson R, Crowell B, Hill B, et al. The effects of L-dopa on the activity of methionine adenosyltransferase: relevance to L-dopa therapy and tolerance. Neurochem Res 1993;18:325–30. Chambers ST. Betaines: their significance for bacteria and the renal tract. Clin Sci 1995;88:25-7 [review]. Charlton CG, Crowell B Jr. Parkinson’s disease-like effects of S-adenosyl-L-methionine: effects of L-dopa. Pharmacol Biochem Behav 1992;43:423–31. Charlton CG, Mack J. Substantia nigra degeneration and tyrosine hydroxylase depletion caused by excess S-adenosylmethionine in the rat brain. Support for an excess methylation hypothesis for parkinsonism. Mol Neurobiol 1994;9:149–61. Cheng H, Gomes-Trolin C, Aquilonius SM, et al. Levels of L-methionine S-adenosyltransferase activity in erythrocytes and concentrations of S-adenosylmethionine and S-adenosylhomocysteine in whole blood of patients with Parkinson’s disease. Exp Neurol 1997;145:580–5. Crowell BG Jr, Benson R, Shockley D, Charlton CG. S-adenosyl-L-methionine decreases motor activity in the rat: similarity to Parkinson’s disease-like symptoms. Behav Neural Biol 1993;59:186–93. Selhub J. Homocysteine metabolism. Annu Rev Nutr 1999;19:217-46 [review]. Homocysteine: Brosnan JT, Jacobs RL, Stead LM, Brosnan ME. Methylation demand: a key determinant of homocysteine metabolism. Acta Biochim Pol. 2004;51(2):405-13. Review. PMID: 15218538 Gahl WA, Bernardini I, Chen S, et al. The effect of oral betaine on vertebral body bone density in pyridoxine-non-responsive homocystinuria. J Inherit Metab Dis 1988;11:291-8. Olthof MR, van Vliet T, Boelsma E, Verhoef P. Low dose betaine supplementation leads to immediate and long term lowering of plasma homocysteine in healthy men and women. J Nutr. 2003 Dec;133(12):4135-8. PMID: 14652361 Olthof MR, Verhoef P. Effects of betaine intake on plasma homocysteine concentrations and consequences for health. Curr Drug Metab. 2005 Feb;6(1):15-22. PMID: 15720203 Schwab U, Torronen A, Toppinen L, Alfthan G, Saarinen M, Aro A, Uusitupa M. Betaine supplementation decreases plasma homocysteine concentrations but does not affect body weight, body composition, or resting energy expenditure in human subjects. Am J Clin Nutr. 2002 Nov;76(5):961-7. PMID: 12399266 Selhub J. Homocysteine metabolism. Annu Rev Nutr 1999;19:217-46 [review]. van Guldener C, Janssen MJ, de Meer K, et al. Effect of folic acid and betaine on fasting and postmethionine-loading plasma homocysteine and methionine levels in chronic haemodialysis patients. J Intern Med 1999;245:175-83. Wendel U, Bremer HJ. Betaine in the treatment of homocystinuria due to 5,10-methylenetetrahydrofolate reductase deficiency. Eur J Pediatr 1984;142:147-50. Wilcken DE, Wilcken B, Dudman NP, Tyrrell PA. Homocystinuria—the effects of betaine in the treatment of patients not responsive to pyridoxine. N Engl J Med 1983;309:448-53. Wilcken DE, Dudman NP, Tyrrell PA. Homocystinuria due to cystathionine beta-synthase deficiency--the effects of betaine treatment in pyridoxine-responsive patients. Metabolism. 1985 Dec;34(12):1115-21. PMID: 3934499 Liver function: Babucke G, Sarre B. Clinical experience with betain citrate. Med Klin 1973;68:1109-13 [in German]. Barak AJ, Beckenhauer HC, Badakhsh S, Tuma DJ. The effect of betaine in reversing alcoholic steatosis. Alcohol Clin Exp Res 1997;21:1100-2. Barak AJ, Beckenhauer HC, Matti J, Tuma DJ. Dietary betaine promotes generation of hepatic S-adenosylmethioine and protects the liver from ethanol-induced fatty infiltration. Alcohol Clin Exp Res 1993;17:552-5. Barak AJ, Beckenhauer HC, Tuma DJ. Betaine, ethanol, and the liver: a review. Alcohol 1996;13:395-8 [review]. PMID: 8836329 Freed WJ. Prevention of strychnine-induced seizures and death by the N-methylated glycine derivatives betaine, dimethylglycine and sarcosine. Pharmacol Biochem Behav. 1985 Apr;22(4):641-3. PMID: 2581277 Junnila M, Barak AJ, Beckenhauer HC, Rahko T. Betaine reduces hepatic lipidosis induced by carbon tetrachloride in Sprague-Dawley rats. Vet Hum Toxicol 1998;40:263-6. Ji C, Kaplowitz N. Betaine decreases hyperhomocysteinemia, endoplasmic reticulum stress, and liver injury in alcohol-fed mice. Gastroenterology. 2003 May;124(5):1488-99. PMID: 12730887 Kettunen H, Tiihonen K, Peuranen S, Saarinen MT, Remus JC. Dietary betaine accumulates in the liver and intestinal tissue and stabilizes the intestinal epithelial structure in healthy and coccidia-infected broiler chicks. Comp Biochem Physiol A Mol Integr Physiol. 2001 Nov;130(4):759-69. PMID: 11691612 Kim SK, Kim YC, Kim YC. Effects of singly administered betaine on hepatotoxicity of chloroform in mice. Food Chem Toxicol 1998;36:655-61. McCarty MF. Co-administration of equimolar doses of betaine may alleviate the hepatotoxic risk associated with niacin therapy. Med Hypotheses. 2000 Sep;55(3):189-94. PMID: 10985907 Murakami T, Nagamura Y, Hirano K. The recovering effect of betaine on carbon tetrachloride-induced liver injury. J Nutr Sci Vitaminol 1998;44:249-55. Poschl G, Stickel F, Wang XD, Seitz HK. Alcohol and cancer: genetic and nutritional aspects. Proc Nutr Soc. 2004 Feb;63(1):65-71. Review. PMID: 15070439 Semmler F. Treatment of liver diseases, especially of fatty liver with betaine citrate. Ther Ggw 1977;116:2113-24 [in German]. Zapadniuk VI, Panteleimonova TN. [Cholagogic effect of trimethylglycine in normal animals of different ages and in experimental atherosclerosis] Biull Eksp Biol Med. 1987 Jul;104(7):30-2. Russian. PMID: 3620644 Autism & Seizures: Rimland B. Seizures, Vitamin B6, DMG, and Sudden Speech. Autism Research Review International. 1996;10(2):1. Roach ES, Carlin L. N,N-dimethylglycine for epilepsy. N Engl J Med. 1982;307:1081-82. Vitamin B6/DMG. Letters to the Editor, Autism Research Interview International. 1994;8(2):6. Immunity: Reap EA, Lawson JW. Stimulation of the immune response by dimethylglycine, a nontoxic metabolite. J Lab Clin Med. Apr1990;115(4):481-6. Safety: Hoorn AJ. Dimethylglycine and chemically related amines tested for mutagenicity under potential nitrosation conditions. Mutat Res. 1989 Apr;222(4):343-50. PMID: 2468082
(https://vitanetonline.com:443/forums/Index.cfm?CFApp=1&Message_ID=988) Introducing... CardioFit Plus Lozenges! 
Date:
September 10, 2005 02:21 PM
Is Your Cardiovascular system Fit? Delicious Homocysteine Lozenges naturally flavored with Orange Creamsicle flavor and REAL orange Juice!
Advanced B-6, B-12, Folic Acid Formula!One Tasty Orange Creamsicle Lozenge Provides:
What is Homocysteine?Homocysteine is an amino acid found in the blood. Healthy homocysteine levels are important markers of normal, healthy cardiac function. Intake of Vitamins B-6, B-12 and folic acid may help provide nutritive support for healthy homocysteine levels. Choline and Trimethylglycine (TMG) act as methyl donors, which can help the body convert homocysteine into methionine.
(https://vitanetonline.com:443/forums/Index.cfm?CFApp=1&Message_ID=777) | ||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||


 We all know that vitamins and other nutrients are essential for a healthy life and if we lack any of these nutrients or vitamins we consume it by supplement. However, most of us do not know that many other substances are also there that work like vitamins, but they are different from Vitamin and TMG or Trimethylglycine is one of these substances that help your body work effectively.